
1.33 Million-Entry Livestock Manure Virome Reframes AMR Risk

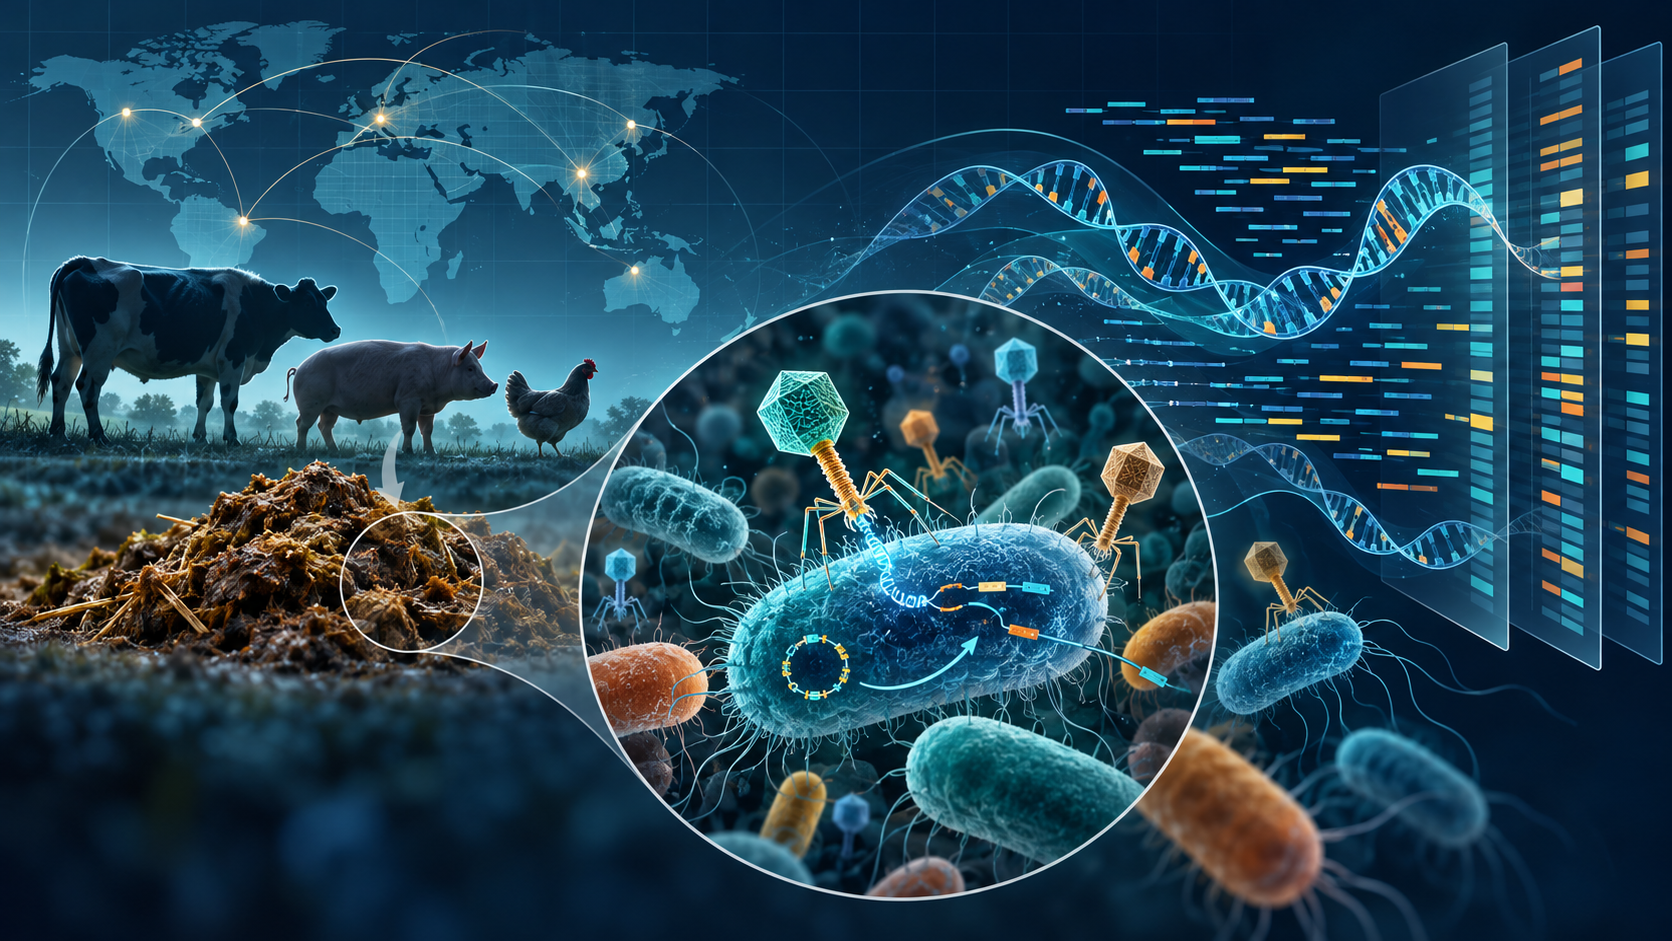
A new global catalogue built from 4,017 livestock-manure metagenomes has assembled 1,330,372 viral operational taxonomic units, creating an unusually large map of the viruses that coexist with bacteria in animal waste. The study’s most consequential result is not that manure viruses commonly carry antibiotic-resistance genes. It is almost the opposite: direct carriage was infrequent, while predicted ecological links between viruses and antibiotic-resistant bacteria were extensive.
The peer-reviewed study, published on 23 October 2026 in Nature Communications, was led by researchers at Northwest A&F University with collaborators in China, South Korea and the United States. The authors call the resource the Livestock Manure Virus Database, or LMVD.
What the 1.33 million entries actually represent
The catalogue’s entries are viral operational taxonomic units, usually shortened to vOTUs. They are sequence-based groupings reconstructed from metagenomic data—not 1.33 million named virus species, confirmed pathogens or disease cases. Most viruses in manure-associated microbial communities are bacteriophages, viruses that infect bacteria rather than livestock or people.
That distinction matters because a database can be enormous without implying a corresponding number of hazards. Its value is analytical: it gives researchers a reference against which new manure samples can be compared, and a framework for asking where particular viruses occur, which bacterial hosts they may infect and how those relationships vary across production systems.
The researchers found that viral communities were strongly geographically restricted. In their modelling, antibiotic use and farming scale had the highest relative predictive importance for viral diversity. Those are associations within a large observational dataset, not proof that changing either factor would produce a specified viral response on an individual farm.
A narrower role for direct resistance-gene carriage
Across the reconstructed viral genomes, the team detected 78 antibiotic-resistance gene subtypes. Those subtypes represented 9.1% of the diversity of resistance genes identified in the wider manure resistome, yet virus-encoded resistance genes were infrequent overall. The result puts an important boundary around one widely discussed pathway: viruses appear unlikely to be a dominant direct container of resistance genes in these samples.
That does not make viruses irrelevant to antimicrobial resistance. The study predicted extensive links between bacteriophages and bacterial hosts. Nearly all antibiotic-resistant bacteria in the dataset were linked to viruses, predominantly temperate phages, and those links were enriched among pathogenic antibiotic-resistant bacteria.
Temperate phages can integrate their genetic material into a bacterial genome and persist as part of the host cell’s biology. Such lysogenic relationships can alter bacterial fitness, competition and gene expression. In this study, however, the links were inferred computationally. They do not demonstrate that a particular phage moved a resistance gene, increased resistance in an animal, or caused a clinical infection.
Why the distinction changes manure-AMR surveillance
Antimicrobial-resistance monitoring often counts resistance genes and profiles bacterial communities. The new catalogue suggests that surveillance may gain explanatory power by also tracking viral–host networks: which phages are associated with which bacteria, whether those relationships persist after manure treatment, and whether temperate phages help particular resistant strains survive or compete.
For farms, manure processors and regulators, this is a research priority rather than a ready-made operating rule. The paper does not compare composting, anaerobic digestion, storage or land-application regimes, and it does not establish that one treatment controls the relevant viral interactions better than another. Those questions now have a much larger reference catalogue against which they can be tested.
The finding also helps separate two risk questions that are sometimes collapsed. One is whether viruses themselves carry resistance genes. The other is whether viruses influence the bacterial populations that carry those genes. The first pathway was uncommon in this dataset; the second appears widespread enough to merit experimental follow-up.
Three findings, three limits
1. Scale: 1,330,372 vOTUs provide a major search and comparison resource. They are not equivalent to 1.33 million named viral species or documented threats.
2. Resistance genes: 78 subtypes occurred in viral genomes, accounting for 9.1% of resistome diversity, but direct viral carriage was infrequent. Diversity is not the same as abundance, transfer rate or health impact.
3. Host links: predicted associations connected nearly all resistant bacteria to viruses, mostly temperate phages. Prediction identifies plausible ecological relationships; it does not establish causation or prove gene transfer.
What researchers and producers should watch next
The decisive next step is experimental validation. Researchers will need cultured systems or carefully designed farm and treatment studies to test whether the predicted phage–host pairs are active, whether lysogeny changes bacterial survival or resistance expression, and how manure handling alters those relationships over time.
Geographic coverage also deserves scrutiny. The paper reports restricted biogeographic patterns, meaning a global reference does not eliminate local sampling gaps. A virus–host relationship seen in one production region may not generalise to a different livestock system, climate, antibiotic-use regime or manure-management practice.
For comparison, a previous Agrilinkage analysis of a controlled broiler study found that chicken age shaped resistance-gene profiles more strongly than the tested feed additives. Together, the studies underline why AMR ecology rarely has a single lever: host age, bacterial community structure, production conditions and viral interactions can all affect the patterns measured in a sample.
Data access and methodology
The authors made the LMVD-associated data and scripts available through a public Zenodo record. Agrilinkage reviewed the paper’s abstract, methods framing, quantitative results, limitations and linked data record. We report virus–host relationships as predicted associations and do not infer clinical transmission, farm-level causality or treatment efficacy from the metagenomic analysis.
Primary source: Jiang et al., “A global livestock virome catalog links viruses with resistome assembly,” Nature Communications (23 October 2026). The article is published under a Creative Commons Attribution 4.0 licence.


Comments